Refinement on Tilt Series
If the tomogram is opened from a tilt series, the alignment of the tilt series can be refined using the splines or molecules.
Coarse Tilt Series Alignment
 New in v1.3.0
API: align_tilt_series
GUI: STA widget > Alignment > Align tilt series
This method uses splines as fiducial markers to align the tilt series. Segments of the splines are represented as a feature-less cylinder, and their projections are used to calculate the cross correlation with the tilt images. Shift, rotation, and tilt angles are optimized at the same time.
Alignment can be improved iteratively. Therefore, tilt series aligned only by IMOD's
tiltxcorr can also be used.
Warning
If filaments (such as microtubules) are oriented toward the same direction, this method is likely to fail. You will usually need to select 3 - 5 filaments that are oriented in different directions.
Template-based Tilt Series Refinement
New in v1.3.0
API: refine_tilt_series
GUI: STA widget > Alignment > Refine tilt series
The subtomograms of all the selected layers are averaged as the reference of all the molecules. The projections of the average placed at the molecules are compared with the tilt images, and the shifts of each tilt image are refined.
List of parameters
- "layers" is the molecules layer(s) used for the refinement. All the layers are averaged together, so they should be of the same structure (such as tubulins of microtubules).
- "size (nm)" is the size of the average. If not given, it tries to use the size of the template image.
- "max shift (nm)" is the maximum shift of the tilt images.
- "grid shape (y, x)" is the number of control points of the local shifts along y
and x of the tilt images.
(1, 1)means a uniform shift of each tilt image. Otherwise, the shifts are refined as a smooth field over each tilt image. - "smoothness" is the weight of the smoothness prior of the local shifts. Larger value is more robust against noise and fewer molecules.
- If "apply CTF" is checked, the CTF is applied to the projections of the average, with the defocus of each molecule in each tilt (changed by its depth in the microscope). The tilt series must have the CTF parameters.
- "bin size" is the bin size of the subtomograms used for the average and the refinement.
- If "update coarse tomogram" is checked, the coarse tomogram for visualization is also reconstructed from the refined tilt series, and shown with the same filter and inversion.
The molecules should be aligned beforehand. align_averaged usually works very well.
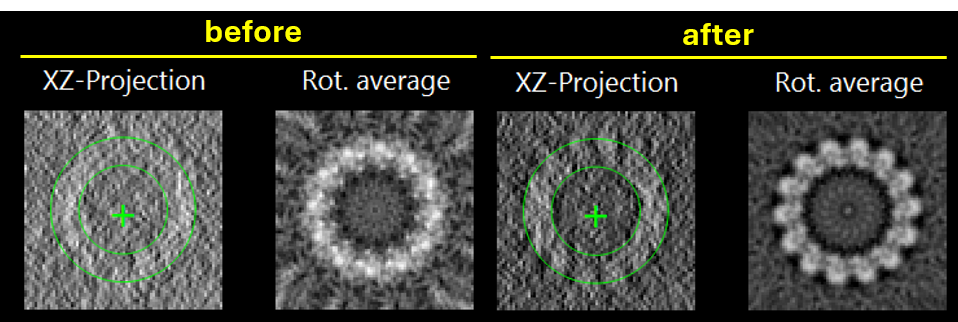
The shifts of the tilt images are shown in the logger. After the refinement, all the subtomograms and the images for analysis are reconstructed from the refined tilt series.
Note
For periodic structures such as microtubules, "max shift (nm)" should be smaller than half of the period (about 2 nm for the 4-nm repeat of tubulin), otherwise the tilt images may be aligned to the neighboring repeat.
The refined tilt series alignment (and the reconstructed coarse tomogram, if checked in
the dialog) are saved in the project directory (tilt_series.json and coarse_tomogram.mrc)
when the project is saved.